
Dr. Yu Chen
ProfessorResearch
5) Iodine/Persulfate-Mediated C–N Bond Formation
Our group has developed metal-free oxidative C–N coupling strategies using iodine reagents and potassium persulfate, which proceed via single-electron-transfer (SET) pathways to enable unconventional bond-forming processes.
In one example, n-Bu₄NI/K₂S₂O₈-mediated transformylation uses aromatic aldehydes as formyl sources to synthesize N-formyl amides. The reaction cleaves a Csp²–Csp² bond in p-anisaldehyde and accepts a broad range of amides (aromatic, aliphatic, and heterocyclic), with gram-scale demonstration. A broader study revealed that electron-donating groups on aromatic aldehydes favor the transformylation pathway, while aliphatic and electron-neutral/deficient aldehydes undergo cross-dehydrogenative C-N coupling via an acyl radical intermediate. Both pathways proceed through SET mechanisms, supported by DFT calculations and radical trapping experiments.
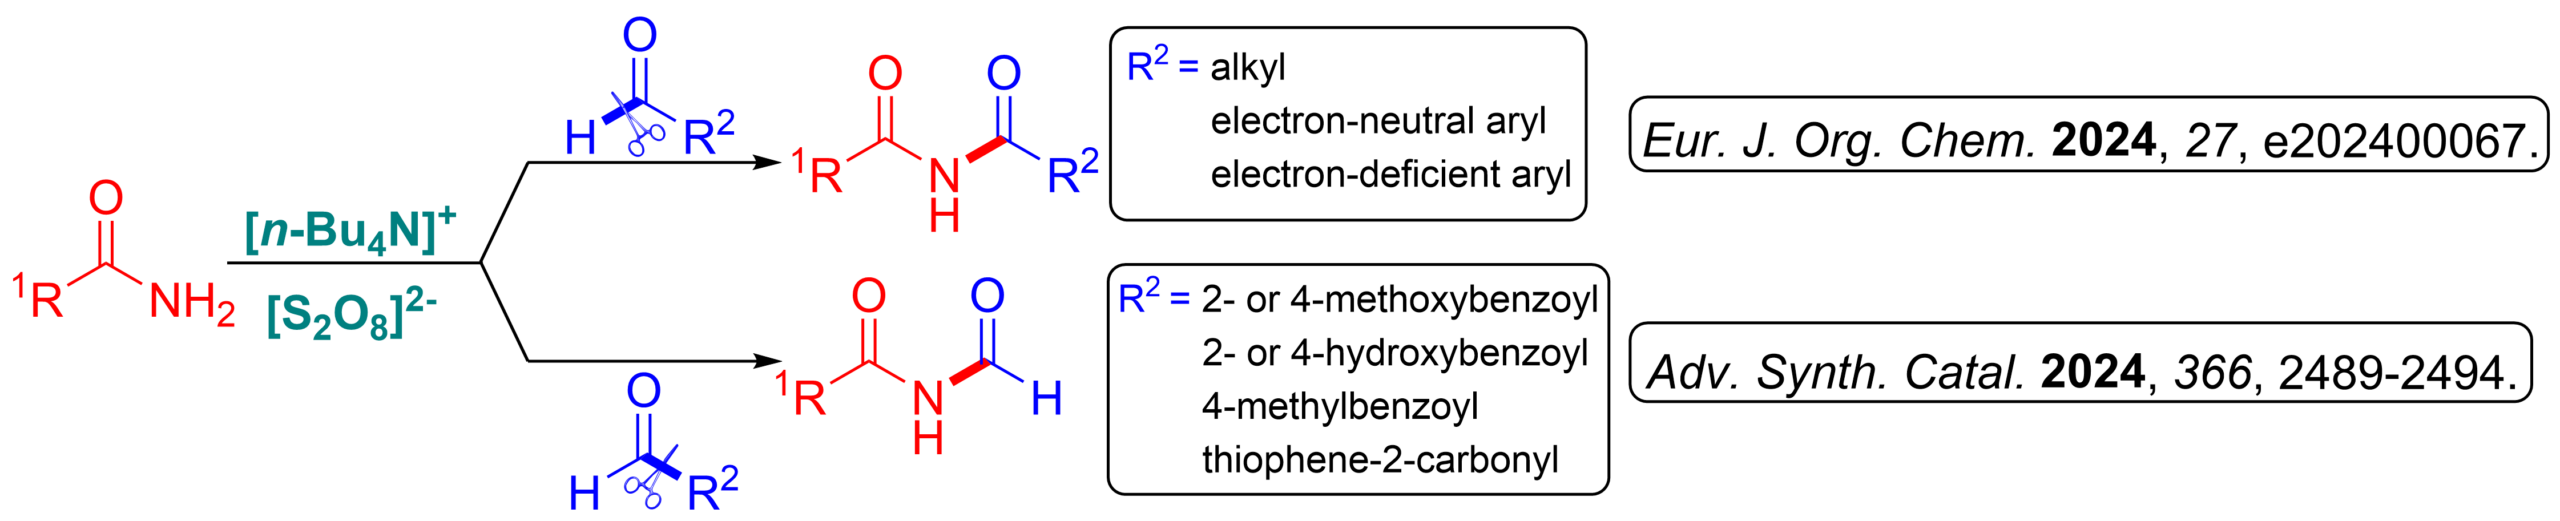 Mechanistic and computational studies revealed an unusual SET pathway involving a phenyl radical cation and an arenium-ion intermediate in the transformylation, demonstrating how iodine/persulfate-mediated radical chemistry can promote C–C bond activation and direct intermediates toward selective C–N bond formation.
More broadly, our work seeks to develop metal-free C–N bond construction by harnessing iodine-derived radical processes, aiming to complement traditional transition-metal catalysis through control of electron-transfer pathways and transient intermediate reactivity.
4) Iron-Catalyzed Reductive Ring Opening
Iron, the second most abundant metal element in the Earth’s crust, represents an attractive alternative to precious metals because of its low cost, abundance, and favorable environmental profile. Iron catalysts can also exhibit distinctive reactivity that complements the chemistry of noble transition metals, making iron catalysis an increasingly important area of synthetic chemistry.
Our group recently developed an iron-catalyzed reductive ring-opening reaction of isoxazoles and isoxazolines through a transfer-hydrogenation pathway. In this transformation, aged N-methyl-2-pyrrolidone serves a dual role as both solvent and hydrogen donor, providing a simple and practical reaction system.
We further leveraged this reductive ring-opening strategy in two iron-catalyzed cascade reactions, enabling the synthesis of both furan and pyrrole derivatives. These studies demonstrate how the unique reactivity of earth-abundant iron can be harnessed to develop efficient cascade processes for the construction and functionalization of heterocyclic molecules.

3) ICl-Mediated Functional Group Interconversion and Electrophilic Cyclization
Iodine-mediated electrophilic reactions provide powerful and versatile approaches for the rapid transformation of unsaturated molecules under mild conditions. Our group has explored iodine monochloride (ICl) as a versatile reagent for both intramolecular electrophilic cyclization and unusual functional group interconversion, developing new strategies for the synthesis of structurally complex and highly functionalized molecules.
Our studies initially focused on ICl-induced intramolecular electrophilic cyclization of alkynes. We developed an ICl-mediated cyclization of 1-([1,1′-biphenyl]-2-yl)alkynones, providing 6-iododibenzocyclohepten-5-ones in high yields. The iodine substituent introduced during cyclization serves as a versatile functional handle for subsequent molecular elaboration, including palladium-catalyzed cross-coupling reactions.
In a complementary direction, our work on ICl-mediated electrophilic cyclization revealed a pronounced substituent-dependent regioselectivity. Incorporation of a para-methoxy substituent on the distal phenyl ring redirected the electrophilic iodocyclization from the expected ortho position to the ipso position of the 1,1′-diphenyl bond, leading to a new family of spiroconjugated compounds. Remarkably, subtle changes in substrate structure allowed the ortho and ipso cyclization pathways to be selectively switched on or off.

During the development of this chemistry, we uncovered an unexpected and highly selective ICl-mediated functional group interconversion. Rather than undergoing the anticipated cyclization, methyl homopropargyl ethers were transformed directly into α-iodo-γ-chloro ketones. This transformation converts a readily accessible alkyne ether into a highly functionalized carbonyl compound while simultaneously installing iodine and chlorine at distinct positions. The reaction accommodates a variety of functional groups and provides a regioselective route to α,γ-dihalo ketones, a valuable class of synthetic building blocks.
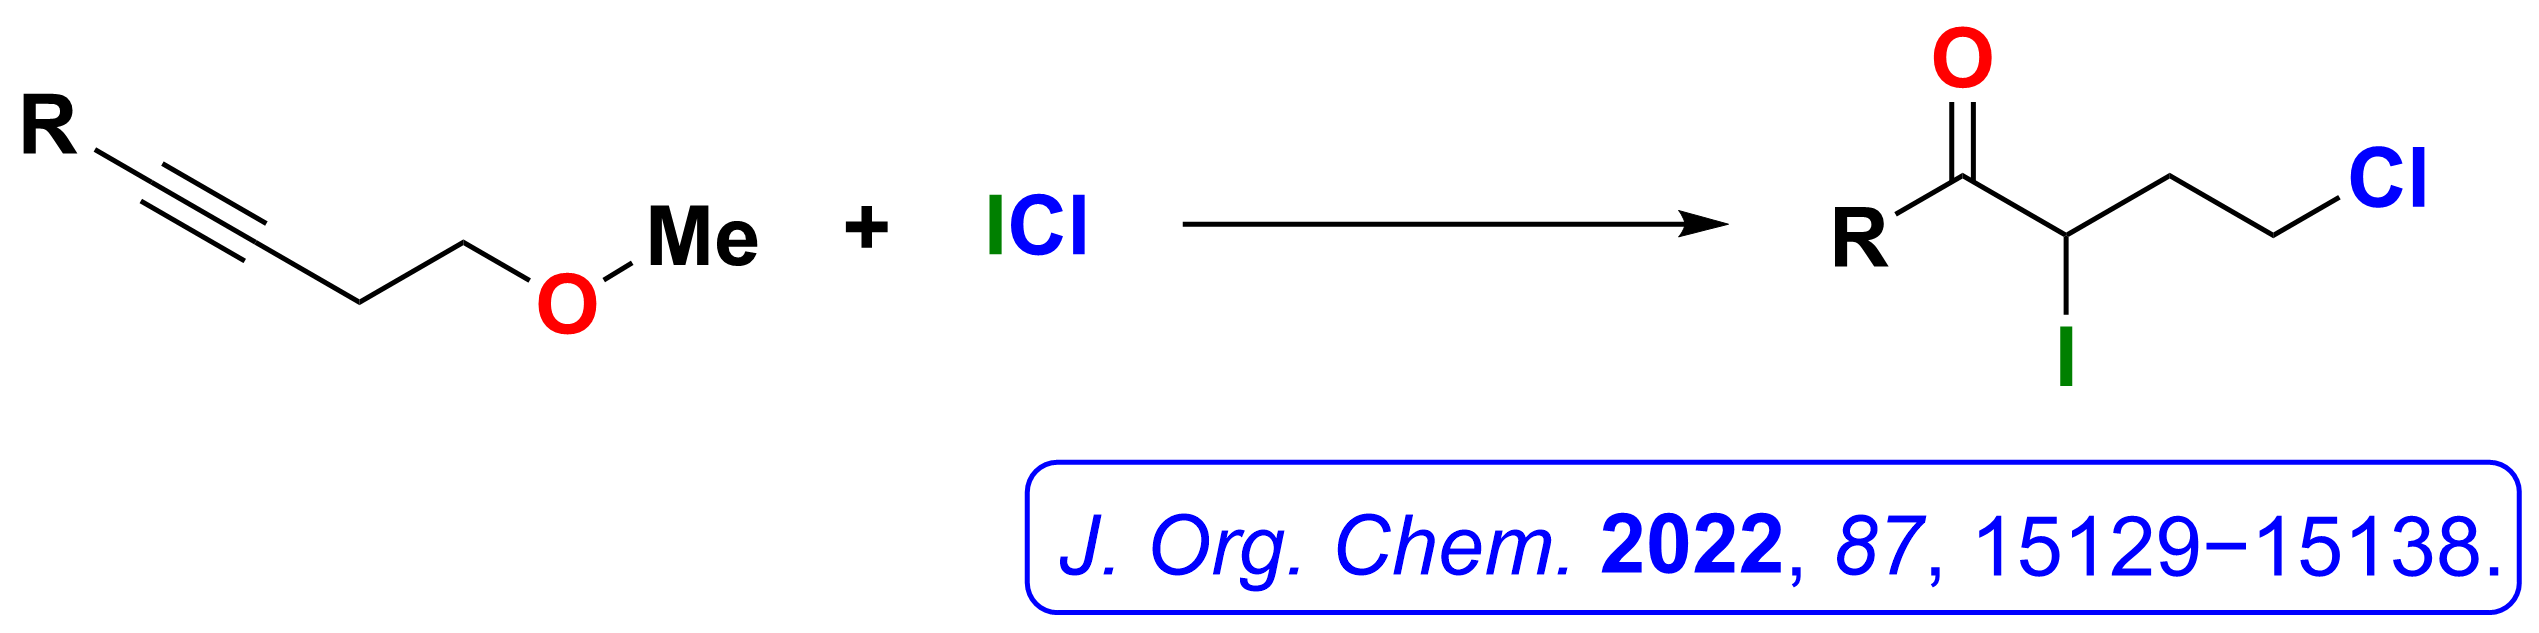
Mechanistic studies revealed that the transformation proceeds through an ICl-induced 5-endo-dig cyclization to form a five-membered oxonium intermediate, followed by chloride-induced ring opening and subsequent demethylation and keto–enol tautomerization. DFT calculations provided insight into the origin of the remarkable chemoselectivity, showing that the ring-opening pathway leading to the α-iodo-γ-chloro ketone is both kinetically and thermodynamically favored over the competing pathway leading to a furan.
Together, these studies demonstrate the versatility of ICl-mediated chemistry for both constructive cyclization and strategic functional group interconversion. By exploiting the electronic properties of substrates and the inherent reactivity of iodine-mediated electrophilic processes, our work provides new approaches to controlling reaction pathways and constructing structurally diverse, highly functionalized molecular architectures.
2) Palladium-Catalyzed Auto-Tandem Chemistry
Palladium-catalyzed cascade and auto-tandem reactions provide powerful strategies for the rapid construction of structurally complex molecules by orchestrating multiple bond-forming events within a single catalytic system. Our group has systematically investigated palladium-catalyzed auto-tandem chemistry, in which a single palladium catalyst promotes sequential, mechanistically distinct transformations to efficiently convert simple substrates into complex molecular architectures.
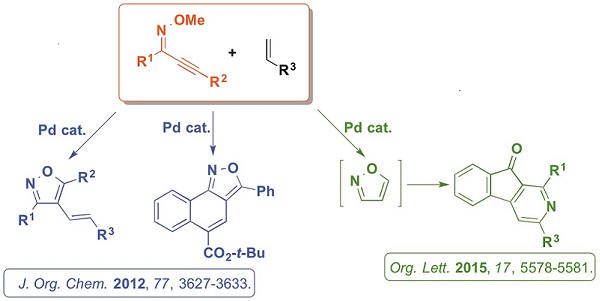
Our early work focused on the synthesis of heterocyclic compounds through palladium-catalyzed cascade reactions. We first developed a palladium-catalyzed cyclization–alkenylation cascade for the synthesis of 3,4,5-trisubstituted isoxazoles. Building on this chemistry, we introduced an ortho-aryl bromide functionality into the oxime substrates, enabling a three-step cascade process that furnished naphthoisoxazoles, a class of compounds that includes well-known estrogen-receptor agonists and antagonists. We subsequently discovered an auto-tandem palladium-catalyzed transformation involving isoxazole ring opening followed by rearrangement of the resulting β-imino ketone intermediates, providing an efficient synthesis of 2-azafluorenones.
More recently, we have extended our auto-tandem catalysis platform to the challenging synthesis of fully substituted, unsymmetrical cyclooctatetraenes (COTs). Unlike conventional COT synthesis, which often relies on the coupling of pre-functionalized and highly symmetrical building blocks, our strategy employs simple alkynone O-methyl oximes and alkynes as starting materials. A single palladium catalytic system orchestrates a sequence of cyclization, sequential double carbopalladation, and C–H activation in one pot, enabling the modular construction of previously difficult-to-access unsymmetrical COT architectures. The method provides a broad range of COT derivatives in good to excellent yields, with up to 95% yield across 17 examples.
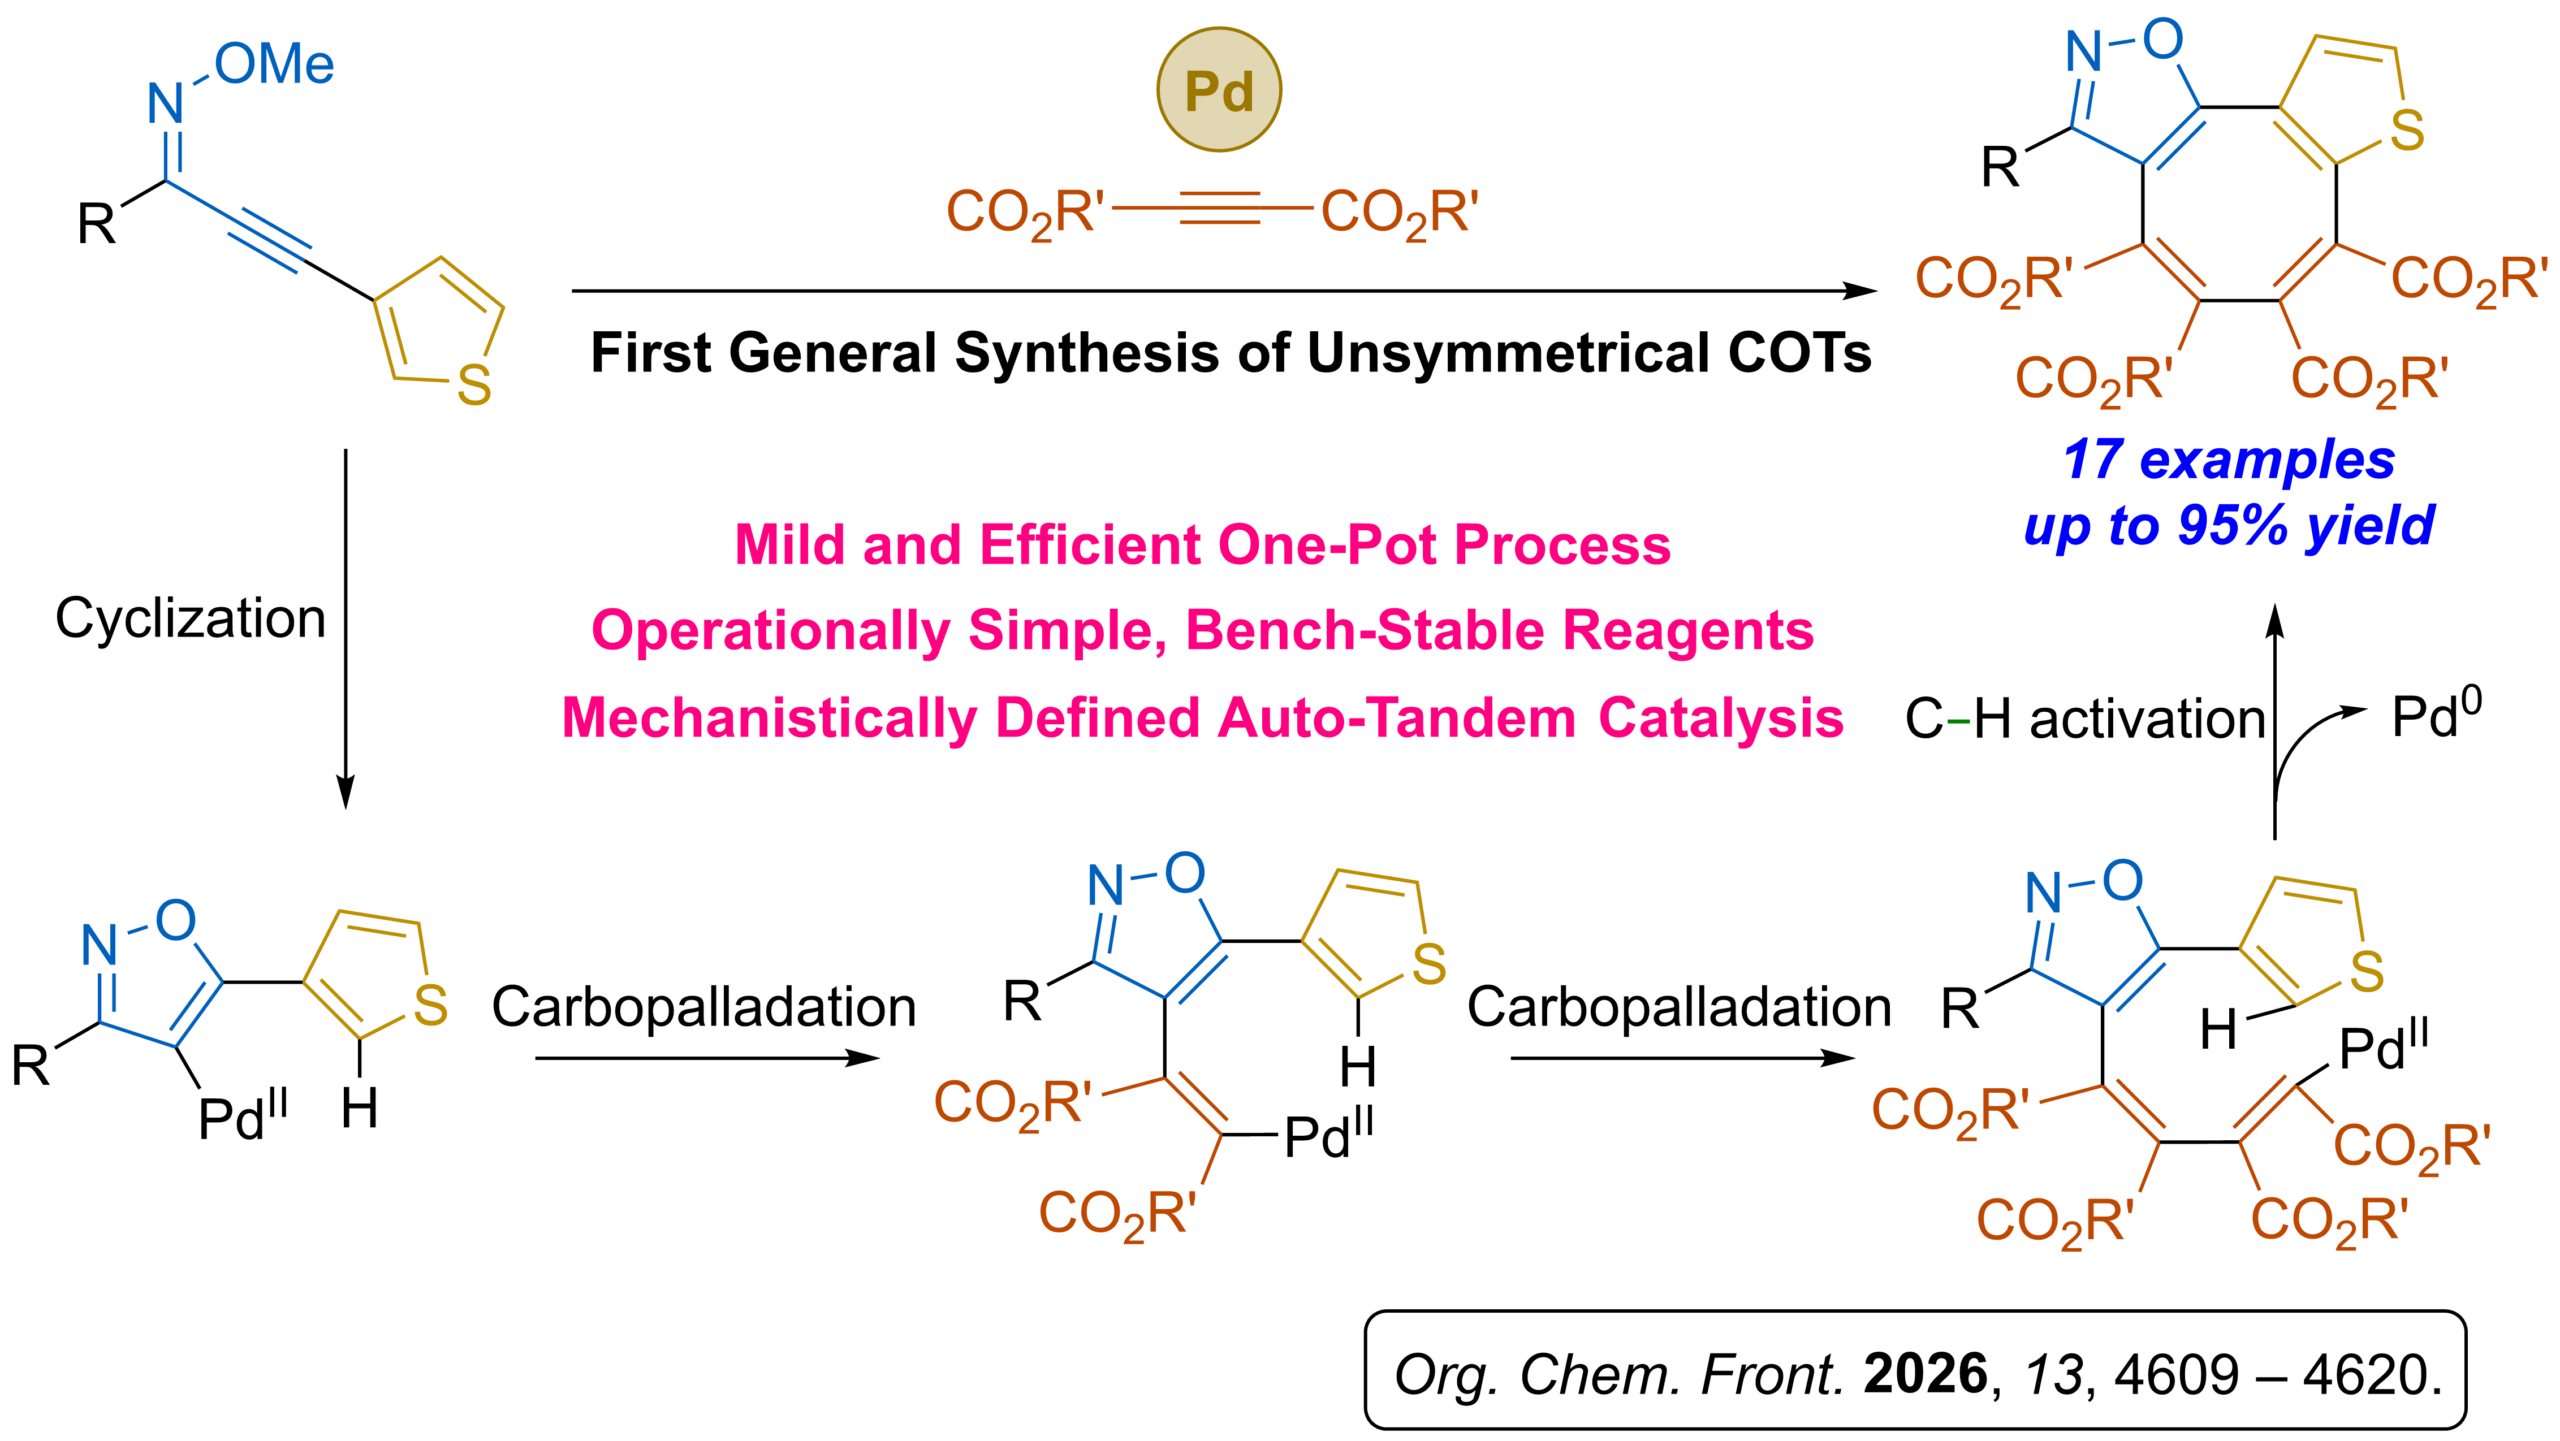
Importantly, our studies combine synthetic development with mechanistic investigation. Experimental studies and DFT calculations revealed the complete catalytic pathway and uncovered the kinetic competition between the second carbopalladation and intramolecular C–H activation that governs product selectivity. The computational analysis further established the origin of regioselectivity and demonstrated how the concentration of the alkyne coupling partner can influence the reaction pathway and product distribution.
Together, these studies establish auto-tandem palladium catalysis as a versatile strategy for the synthesis of complex molecular frameworks. Beyond individual transformations, our work seeks to develop general principles for programming multiple catalytic events within a single reaction manifold, providing new approaches to the efficient construction of structurally and electronically diverse molecules.
1) Nitrogen-Containing Heterocycles from Ammonium Acetate
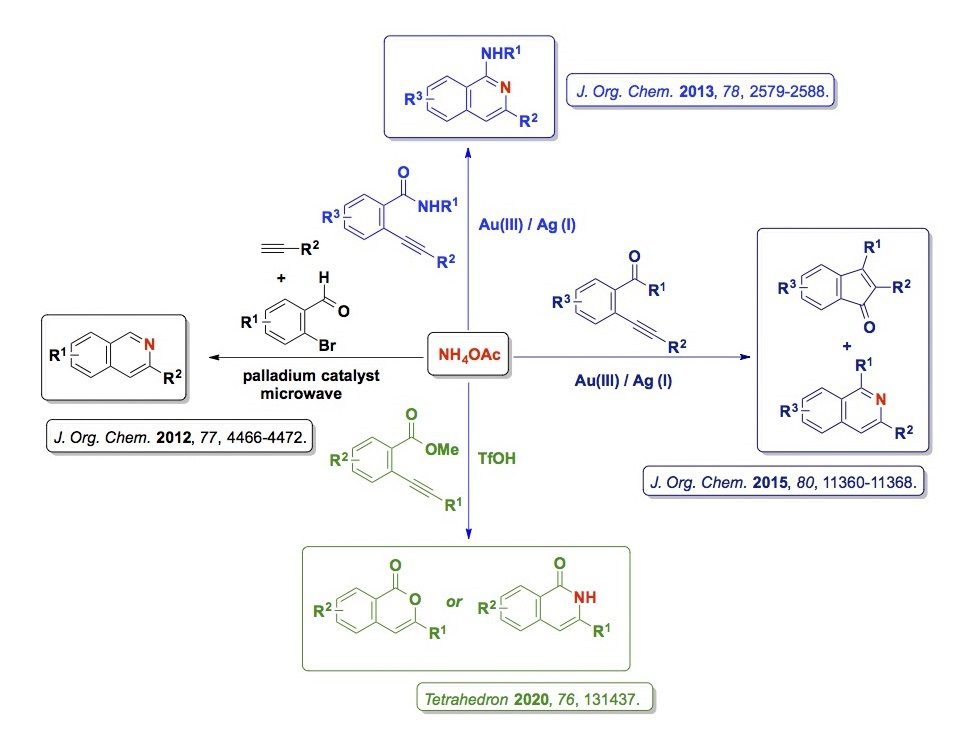
Nitrogen-containing heterocycles are ubiquitous in natural products, pharmaceuticals, and functional materials. Inspired by our interests in late-transition-metal catalysis and heterocyclic chemistry, our group has developed efficient strategies that employ ammonium acetate (NH₄OAc) as a readily available ammonia source for the imination step in transition-metal-catalyzed cascade and one-pot reactions.
Our group first reported a palladium-catalyzed, microwave-assisted one-pot synthesis of isoquinolines. This convenient protocol was subsequently extended to the synthesis of furopyridines and thienopyridines and was highlighted in Synfacts. We further developed gold-catalyzed domino reactions for the synthesis of isoquinolines, 1-aminoisoquinolines, and indenones, as well as a triflic-acid-mediated approach to isoquinolinones.
These studies demonstrate the versatility of ammonium acetate as a practical nitrogen source and provide streamlined approaches to diverse nitrogen-containing heterocyclic scaffolds.